Cystic Fibrosis: Causes, Symptoms, Diagnosis, Treatment
Author: Disabled World (DW)
Updated/Revised Date: 2 Nov 2026
Table of Contents:
Synopsis - Definition - About This Section - FAQs - Publications - Subtopics
Synopsis
An overview of cystic fibrosis, an inherited disorder affecting the lungs and glands, covering its causes, symptoms, diagnosis and available treatments.
At a Glance
- 1 - All U.S. states have screened newborns for cystic fibrosis since 2010.
- 2 - A sweat chloride level of 60 mmol/L or higher indicates cystic fibrosis.
- 3 - Nearly 40,000 children and adults in the United States are living with cystic fibrosis.
- 4 - Half of babies born with cystic fibrosis in 2023 are expected to reach age 68 or older.
Topic Definition
- Cystic Fibrosis
Cystic fibrosis, also known as mucoviscidosis, is an inherited genetic disorder that mainly affects the lungs but also involves the pancreas, liver, kidneys, and intestines. It is caused by a fault in the CFTR gene, which normally helps regulate the movement of salt and water in and out of cells; when this process breaks down, the body produces unusually thick, sticky mucus that clogs the airways and blocks ducts in the digestive organs. The result is a range of long term problems, including difficulty breathing, repeated lung infections, poor growth, fatty stools, and, in most men, infertility, though the severity varies widely from one person to another. Cystic fibrosis passes to a child only when both parents carry the recessive gene, and although there is currently no cure, modern treatments such as airway clearance, antibiotics, enzyme supplements, and careful nutrition allow many people with the condition to live far longer and more active lives than in the past.
Overview
Defining Cystic Fibrosis
Cystic fibrosis (CF), also known as mucoviscidosis, is a genetic disorder that changes a protein in the body, making mucus thick and sticky. It affects mostly the lungs but also the pancreas, liver, kidneys, and intestines. Long-term problems include trouble breathing and coughing up mucus because of frequent lung infections. Other symptoms include sinus infections, poor growth, fatty stools, clubbing of the fingers and toes, and infertility in most men. Symptoms vary from person to person.
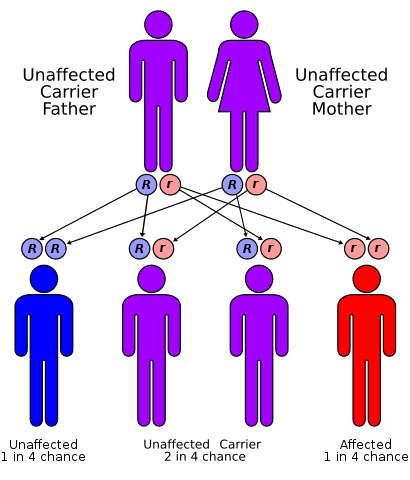
Cystic fibrosis is inherited in an autosomal recessive pattern. A child gets CF only if both parents pass on a faulty copy of the gene. Parents who each carry one copy usually have no symptoms. When both parents are carriers, each child has a 25 percent chance of having CF, a 50 percent chance of being a carrier without CF, and a 25 percent chance of not having the gene at all.
CF is most common in white people in the United States, affecting 1 in 2,500 to 3,500 white newborns. It is less common in other groups, affecting about 1 in 17,000 African Americans and 1 in 31,000 Asian Americans [MedlinePlus Genetics, n.d.].
Cystic fibrosis used to cause death in childhood, but survival has improved greatly thanks to newborn screening, new medicines, better nutrition, and lung transplants. Nearly 40,000 children and adults in the United States, and more than 100,000 worldwide, are living with CF. Children born with CF between 2019 and 2023 are expected to live an average of 61 years, and half of babies born with CF in 2023 are expected to reach age 68 or older [National Heart, Lung, and Blood Institute, 2024]. Lung disease remains the most serious and common complication.
Causes of Cystic Fibrosis
In 1989, researchers found the gene that causes cystic fibrosis, called CFTR, and its most common mutation, known as Delta F508 (F508del). The CFTR gene makes a protein, the cystic fibrosis transmembrane conductance regulator, that works as a channel moving chloride, a part of salt, into and out of cells. The flow of chloride helps control the movement of water in tissues, which is needed to make thin, freely flowing mucus. In CF, the CFTR protein is missing, does not reach the cell surface, or does not work properly. As a result, mucus becomes thick and sticky and builds up in the lungs and digestive organs, and sweat becomes very salty. Many different CFTR mutations have been found, and which ones a person has can affect how severe their CF is and which medicines will work for them.
Symptoms of Cystic Fibrosis
Symptoms vary. For some people they appear soon after birth, while for others they may not appear for months or even years. In infants and babies, signs can include salty-tasting skin, poor weight gain, a blocked intestine at birth (meconium ileus), and pale, bulky, greasy, foul-smelling stools, as well as gas, a swollen belly, and a persistent cough or wheezing.

Children and adults with CF may have a constant cough with thick mucus, wheezing, shortness of breath, frequent lung infections such as pneumonia and bronchitis, and poor growth or weight loss despite a good appetite. Other problems can include:
- Clubbing, a rounding and widening of the fingertips and toes
- Sinus infections and nasal polyps
- A collapsed lung (pneumothorax) or coughing up blood
- Problems with the pancreas, gallbladder, and liver, including CF-related diabetes
- Delayed puberty
- Infertility in most men, and reduced fertility in some women
Diagnosing Cystic Fibrosis
All U.S. states have screened newborns for cystic fibrosis since 2010. The newborn screen measures immunoreactive trypsinogen (IRT), a protein made by the pancreas, in a blood sample taken a few days after birth; high levels may point to CF and are followed by more testing.
The sweat test is the standard test to confirm CF. A chemical called pilocarpine and a mild electric current are used to make the skin on the forearm sweat, and the amount of chloride in the sweat is measured. A chloride level of 60 mmol/L or higher indicates cystic fibrosis. Before birth, CF can be diagnosed with genetic testing of a sample taken by chorionic villus sampling or amniocentesis. Other tests, such as lung function tests, chest X-rays, stool tests, and sputum (phlegm) cultures, help show how CF is affecting the body.
Genetic and Carrier Testing
Genetic testing on a blood sample or a swab from the inside of the cheek can show which CFTR mutations a person has. Carrier testing is often offered to people who are pregnant or planning a pregnancy, and to their partners. Brothers, sisters, and other relatives of a person with CF may also choose to be tested, and genetic counseling can help families understand the results.
Treating Cystic Fibrosis
There is no cure for cystic fibrosis yet, but treatment can improve quality of life and help people live longer. CFTR modulator medicines treat the faulty protein itself: elexacaftor, tezacaftor, and ivacaftor (Trikafta), approved by the U.S. Food and Drug Administration (FDA) in 2019, can be used by most people with CF, and vanzacaftor, tezacaftor, and deutivacaftor (Alyftrek) was approved in December 2024. Care is usually given by a team at a specialized CF center.
Lung Treatments
- Airway clearance: chest physical therapy, in which the chest and back are clapped or vibrated, breathing techniques such as autogenic drainage, handheld devices such as positive expiratory pressure (PEP) and oscillating devices, and inflatable vests that shake the chest at high frequency, all help loosen and clear mucus.
- Inhaled medicines: dornase alfa (Pulmozyme), given with a nebulizer, thins mucus, and inhaled hypertonic saline and bronchodilators can help open the airways.
- Antibiotics: lung infections are treated with antibiotics given by mouth, by vein, or inhaled, such as inhaled tobramycin or aztreonam.
- Anti-inflammatory medicines: some people take high-dose ibuprofen or other medicines to reduce airway inflammation.
- Lung transplant: people with severe lung disease that no longer responds to treatment may be offered a lung transplant.
Digestive and Nutrition Treatments
Most people with CF need a high-calorie diet that is high in protein and fat, along with pancreatic enzyme capsules taken with meals and snacks to help digest food, and vitamins A, D, E, and K. Gallstones may be treated by removing the gallbladder, usually with laparoscopic surgery, and liver disease may be treated with ursodeoxycholic acid (ursodiol).
Fertility
Most men with CF are infertile because the tubes that carry sperm are blocked or missing, but many can have biological children with sperm retrieval and in vitro fertilization with intracytoplasmic sperm injection (ICSI). Many women with CF can become pregnant, but pregnancy needs careful planning and medical care, especially for women with serious lung disease.
Frequently Asked Questions
Is cystic fibrosis contagious?
No, cystic fibrosis is not contagious, since it is an inherited genetic condition passed on only when a child receives a faulty gene from both parents.
What is the life expectancy for someone with cystic fibrosis?
Life expectancy has improved greatly with modern care, and many people with cystic fibrosis now live into their forties, fifties, or beyond.
How is carrier status for cystic fibrosis detected?
A simple genetic test on a saliva or blood sample can show whether a person carries the cystic fibrosis gene, which is useful for family planning.
Can people with cystic fibrosis have children?
Yes, with assisted reproductive techniques many men can father children, and many women can conceive, though pregnancy needs careful medical supervision.
What is chest physiotherapy for cystic fibrosis?
It is a set of techniques and devices that loosen and clear thick mucus from the lungs to reduce infection and help breathing.
Does diet matter for people with cystic fibrosis?
Yes, a high calorie, nutrient rich diet along with enzyme and vitamin supplements helps counter poor nutrient absorption and supports growth and energy.
Why do people with cystic fibrosis have salty skin?
The faulty CFTR protein disrupts salt balance so more salt is lost through sweat, giving the skin a noticeably salty taste.
What are CFTR modulator drugs?
CFTR modulators are medicines that help the faulty CFTR protein work better, targeting the cause of cystic fibrosis rather than its symptoms. Trikafta, approved in 2019, can be used by most people with the disease and has greatly improved lung function and life expectancy.
References
- MedlinePlus Genetics. (n.d.). Cystic fibrosis. National Library of Medicine.
- National Heart, Lung, and Blood Institute. (2024). What is cystic fibrosis?
Curated and edited by Ian C. Langtree, Founder and Editor-in-Chief, Disabled World. This section is maintained by the Disabled World editorial team. Last updated: